Phenylketonuria: Symptoms, Causes, Treatment and Suitable Diet Plan
Smart Voice
AITable of Contents 5
Comprehensive overview of phenylketonuria: a genetically inherited metabolic disorder
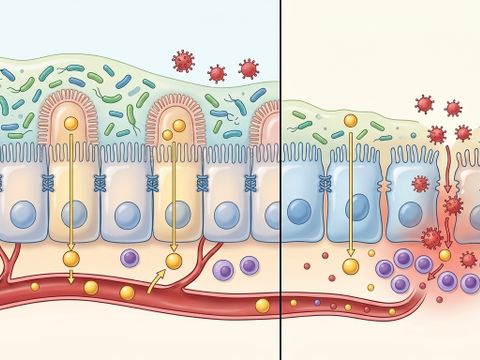
Phenylketonuria represents an inherited metabolic disorder of genetic origin, which is why mandatory newborn screening programs systematically test for its presence shortly after birth. Individuals affected by this condition exhibit an enzymatic deficiency that impairs the body’s ability to convert phenylalanine—an essential exogenous amino acid—into tyrosine, thereby disrupting the biochemical balance between these two compounds. A critical challenge arises from the ubiquity of phenylalanine, which, as a protein constituent, is found in virtually all food products, particularly those of animal origin, including meat, dairy, and eggs. As a result, patients diagnosed with phenylketonuria must adhere to a lifelong, highly restrictive low-phenylalanine diet to prevent severe neurological and developmental complications.
Phenylketonuria: Diagnostic Symptoms and Long-Term Complications of an Untreated Metabolic Disorder
The genetic basis of phenylketonuria: an enzymatic mutation on chromosome 12
Management of phenylketonuria: dietary interventions and their role in preventing neurocognitive impairment
Phenylketonuria dietary management: Essential nutritional guidelines for individuals with phenylalanine metabolism disorders
Want to apply these guidelines in your daily diet?
Our smart diet generator creates a tailored meal plan in 3 minutes based on the recommendations in this article.
Tags

Literatura: Jarochowicz S., Mazur A., Fenyloketonuria – choroba metaboliczna, uwarunkowana genetycznie, w: Przegląd medyczny Uniwersytetu Rzeszowskiego, 2007. Milupa, Fenuloketoniura: Dzieci w 1. roku życia, zasady postępowania dietycznego – poradnik, 2012.
Diets Premium Expert
Diets Premium team expert. Specializes in nutrition plans, strength sports, and training plans.